FDA Q-Sub Question Matrix Template and ISO 13485 to 21 CFR Crosswalk: Two Workbooks That Do the Work
How to build an FDA Q-Sub question matrix that follows FDA's 2025 guidance (7–10 questions, up to four topics) and an ISO 13485 to 21 CFR crosswalk for the QMSR, with example rows, common mistakes and ready-made Excel workbooks.
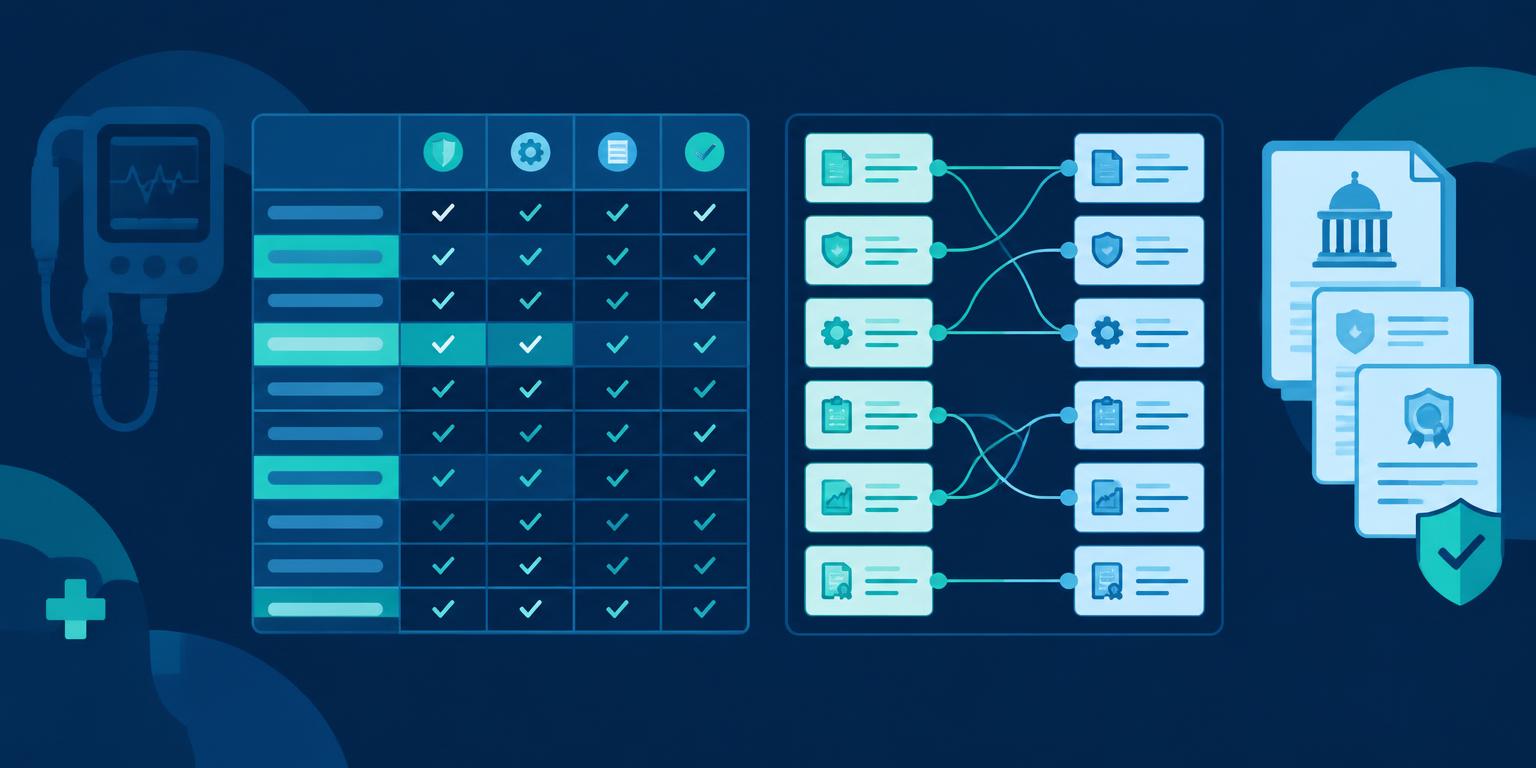
Two searches tend to show up in the same week: "FDA Q-Sub question matrix template" and "ISO 13485 to 21 CFR crosswalk." That's no coincidence. Teams getting ready to ask FDA for feedback are often also moving their quality system onto FDA's Quality Management System Regulation (QMSR), which took effect February 2, 2026. Both jobs come down to the same discipline: put each question or requirement in its own row, name an owner and attach the evidence.
This guide explains how to build both tools, shows what a good row looks like, and points to the Cahir MedTech workbooks that come with the structure already in place.
Part 1: The FDA Q-Sub question matrix
What FDA actually asks for
FDA's final guidance, Requests for Feedback and Meetings for Medical Device Submissions: The Q-Submission Program (issued May 29, 2025), asks submitters for specific questions backed by enough information for FDA to give useful feedback. Three points from the guidance shape the matrix:
- Size. FDA says the most effective Pre-Subs "typically have no more than 7-10 questions (including sub-questions)," split across "no more than four substantial topics."
- Proposals, not open questions. FDA cannot design study plans for you. To get protocol feedback, submit a proposed outline with a rationale and specific questions about it.
- Minutes are yours to write. For Pre-Sub meetings, the submitter drafts the minutes and submits them to FDA as an amendment within 15 days of the meeting.
FDA does not publish an official question-matrix form. The matrix is a planning tool your team builds to meet these expectations.
Illustrative example row
This row is a made-up example, not a real FDA interaction:
- Topic: Analytical performance
- Question: "Does FDA agree that the proposed 120-sample method comparison, using the acceptance criteria in Section 4.2, is sufficient to support the planned 510(k)?"
- Proposed approach: A method comparison against the predicate across three sites, with pre-specified acceptance criteria.
- Follow-up: If FDA asks for more low-concentration samples, the study lead updates the protocol before enrollment.
Common mistakes
1. Too many questions. More than 7–10 weakens the feedback on each one, and FDA may ask you to cut topics. 2. Open-ended asks such as "What testing does FDA require?" with no proposal attached. 3. Questions that depend on each other. When one answer changes another question, FDA's feedback gets harder to apply. 4. No follow-through. Feedback recorded in a slide deck never makes it into the protocol or the design history.
The workbook
Watch the open Q-Sub question-matrix video walkthrough to see the method; no registration needed. The FDA Q-Sub Master Pack ($499, one-time per company workspace) includes a fourteen-item Q-Sub readiness baseline and type selector, the question matrix, a briefing document builder, a feedback tracker and a timeline planner back-planned from protocol lock. The Q-Sub companion guide explains each tab.
Part 2: The ISO 13485 to 21 CFR crosswalk
Why certification isn't enough
Under the QMSR, 21 CFR Part 820 incorporates ISO 13485:2016 by reference (820.10) and adds US-specific requirements, for example records (820.35) and labeling and packaging controls (820.45). ISO 13485 refers again and again to "applicable regulatory requirements" but never lists them. A crosswalk is where you name them. Our November 6 article on mapping 21 CFR to ISO 13485 covers the full method; here is the working structure.
Cropped crosswalk sample
Swipe horizontally to see all columns.
| ISO clause | US requirement | Evidence example |
|---|---|---|
| 8.2.3 Reporting | 21 CFR Part 803 | Documented MDR evaluation |
This cropped example is not the full schema. The complete crosswalk and tracking structure are included in the paid kit.
Rows most teams need
- Part 820: QMSR additions to ISO 13485 (820.10, 820.35, 820.45)
- Part 803: medical device reporting, mapped to clause 8.2.3
- Part 806: reports of corrections and removals, mapped to 8.3 and advisory notices
- Part 807: establishment registration, device listing and premarket notification
- Part 801 and Part 830: labeling and unique device identification
Common mistakes
1. Treating the ISO certificate as proof of compliance with FDA requirements. 2. Mapping clauses with no owning procedure or evidence. 3. Forgetting the parts that sit outside Part 820, such as 803, 806 and 830. 4. Using an unauthorized or outdated copy of the standard.
The workbook
The 21 CFR–ISO 13485 Mapping Kit ($199, one-time) includes the crosswalk from ISO 13485 "regulatory requirements" clauses to 21 CFR Parts, a reverse map of US-only additions (MDR, 806, 807, UDI, Part 11), a gap action plan, a procedure register and an inspection checklist. Read the companion guide first.
Templates vs. static standards PDFs
Standards publications, such as those sold by AAMI and ISO, define the requirements, and you still need an authorized copy of any standard that applies to you. A PDF tells you what's expected; a workbook helps your team do it: assign owners and dates, score open gaps, record evidence and sign off, then track the same items together in your company workspace. See how all of our kits compare on the toolkit overview. Unfamiliar terms are defined in the MedTech regulatory glossary.
Frequently asked questions
Does FDA provide an official Q-Sub question template?
No. FDA's Q-Submission guidance describes what a request should contain and recommends specific questions with proposed approaches, but it doesn't publish a question-matrix form. Teams build their own matrix or use a structured workbook.
How many questions should a Pre-Sub include?
FDA's May 2025 guidance says the most effective Pre-Subs typically have no more than 7–10 questions, including sub-questions, across no more than four substantial topics. Lower-priority topics can go into a later Pre-Sub.
Is ISO 13485 certification enough for FDA under the QMSR?
No. Part 820 incorporates ISO 13485:2016 but adds US-specific requirements, and other parts such as 803, 806, 807 and 830 still apply. A crosswalk shows where each one is handled in your quality system.
Do the workbooks replace the ISO 13485 standard?
No. They don't reproduce the standard. Buy an authorized copy, then use the crosswalk to plan, assign and evidence the work.
Does using these workbooks guarantee FDA clearance or a clean inspection?
No. They are planning and coordination tools. FDA's decisions depend on your device, your data and the current requirements, so use qualified regulatory review.
Primary sources
- FDA Q-Submission Program final guidance (May 29, 2025): question count, topics and meeting minutes
- FDA QMSR overview: effective February 2, 2026
- eCFR: Part 820, Part 803, Part 806, Part 807, Part 801, Part 830
- ISO 13485:2016 (ISO)
This article is educational guidance, not legal advice or an FDA-approved template. Publication date: November 9, 2026.
New to a term? See the MedTech regulatory glossary for plain-English definitions.
Follow MedTech Insights
New articles on FDA 510(k) and De Novo pathways, CE Mark and EU MDR, and device reimbursement — sent to your inbox as they publish.