FDA 510(k) Third-Party Review Program: Is It Right for Your Device?
The Third Party Review Program lets an FDA-recognised organisation review your 510(k) first. Here is who is eligible, what it costs, where it saves time, and when it is the wrong choice.
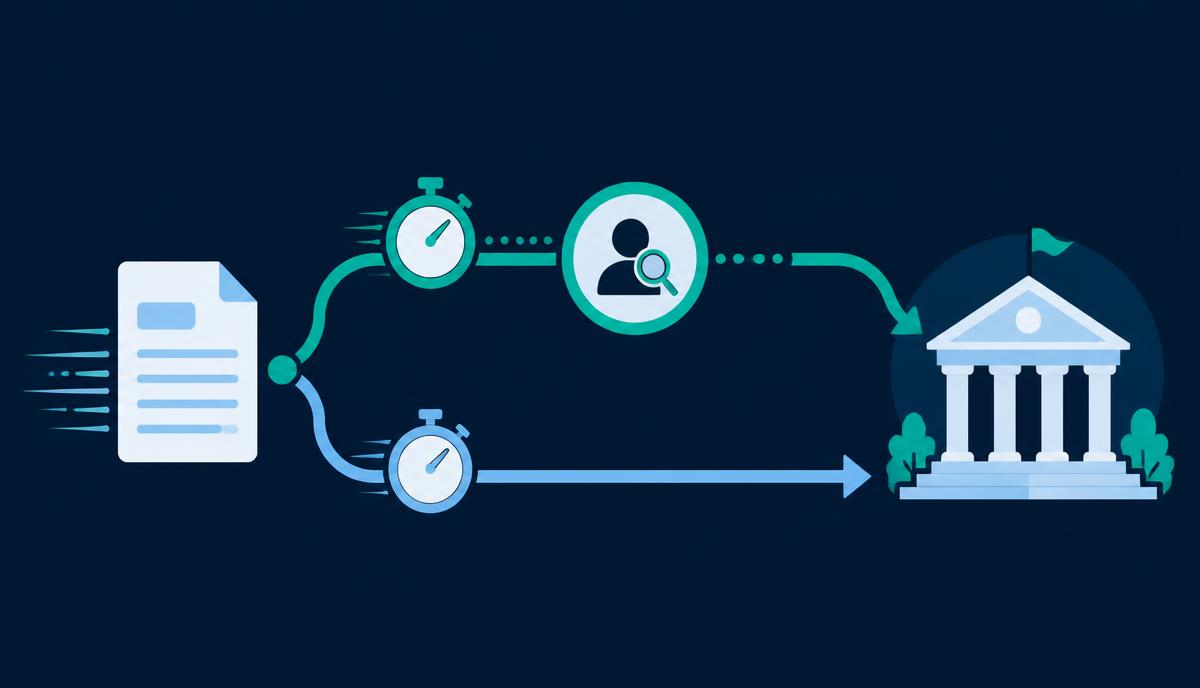
Not every 510(k) has to go to FDA first. Under the Third Party Review Program, an FDA-recognised review organisation can assess your submission and send FDA a recommendation, and FDA then issues the final decision on a compressed statutory clock.
It is a genuinely useful route for the right device, and a waste of money for the wrong one. Here is how to tell which you have.
What the programme actually is
FDA recognises independent organisations — accredited persons — to conduct the primary review of certain 510(k) submissions. You submit to the review organisation rather than to FDA. The organisation reviews the submission, interacts with you on deficiencies, then forwards its recommendation and the full submission to FDA.
FDA makes the final substantial equivalence determination. The programme does not delegate the decision; it delegates the review work. Background on how 510(k) fits alongside De Novo and PMA is in our FDA regulatory pathways guide.
Eligibility is device-specific, not company-specific
This is the first filter, and it is binary. FDA publishes the list of device types eligible for third-party review, organised by product code, and each recognised review organisation is itself recognised only for a defined scope of device types.
Two checks, in order: is my product code on the eligible list, and is there a recognised organisation whose scope includes it? If either answer is no, the programme is not available to you, regardless of how straightforward your device is.
Broadly, eligibility skews to lower-risk, well-characterised Class II devices. Devices requiring clinical data, devices that are implantable or life-sustaining, and anything raising novel safety questions are generally excluded. DevicePath handles exactly this classification and product-code triage.
What you gain
Speed is the main argument. Once the review organisation forwards its recommendation, FDA operates on a short statutory review period, substantially shorter than a standard 510(k) review cycle.
You also avoid the FDA 510(k) user fee. Submissions reviewed under the programme are not subject to the standard 510(k) fee — relevant when the standard fee runs tens of thousands of dollars. Our post on De Novo classification fees explains how the fee tiers and small-business reductions work across submission types.
The third gain is less discussed: interaction. Review organisations are typically more available for dialogue during review than an FDA reviewer with a large queue. For a first-time manufacturer, that access can be worth more than the calendar saving.
What it costs
You pay the review organisation directly, on commercial terms it sets. There is no published fee schedule because there is no single fee.
The honest comparison is the organisation's fee against the FDA user fee you avoid, plus the value of the time saved. For a small business already eligible for a reduced FDA fee, the arithmetic is much tighter than it looks in a vendor pitch, because the fee being avoided is already discounted.
Where it goes wrong
Three failure patterns recur.
The submission was not ready. A review organisation does not lower the bar. A thin predicate comparison or incomplete testing produces the same deficiencies faster, and you have now paid twice for the privilege. If your predicate story is uncertain, read our post on what happens when you do not have a predicate device before choosing a route.
The device was borderline. If your device sits at the edge of its eligible product code, or has a feature that raises a new question of safety, the review organisation may decline it or FDA may disagree with the recommendation. Either way you lose the time you were trying to save.
The scope was wrong. Review organisation recognitions change. Confirm current scope against FDA's published list at the time you plan to submit, not from a list you saved last year.
Where it works well
The programme earns its keep when the device is squarely within an eligible, well-understood product code, the predicate is clean and current, the testing package is complete and standards-based, no clinical data is required, and you want predictability more than you want agency dialogue about a novel feature.
That describes a lot of Class II devices, and teams overlook it because they assume FDA is the only door.
A practical decision sequence
Confirm your product code and its eligibility against FDA's published device list.
Identify recognised review organisations whose current scope covers that code, and check FDA's published performance and accreditation status information for them.
Quote the review fee and compare it against the FDA fee you would actually pay, including small-business reductions.
Assess submission readiness honestly. If a pre-submission meeting would still be useful, do that first — the Q-Submission route and third-party review are not mutually exclusive.
Decide on time value, not fee value. The programme's real product is a shorter, more predictable path for a device that was already going to clear.
Beyond the US
Third-party review has a conceptual cousin in Europe, where notified bodies perform conformity assessment as a matter of course — though the EU model is far broader and the certificate structure is different. Our EU MDR and UKCA compliance guide covers that, and our market access guide covers what comes after clearance in either market.
Mapping submission route, review option and market sequencing together is what MedTech Compass is built for.
Sources
1. FDA, 510(k) Third Party Review Program — https://www.fda.gov/medical-devices/premarket-submissions-selecting-and-preparing-correct-submission/510k-third-party-review-program 2. FDA, How to Submit a 510(k) Through the Third Party Review Program — https://www.fda.gov/medical-devices/510k-third-party-review-program/how-submit-510k-through-third-party-review-program 3. FDA, 510(k) Third Party Performance Metrics and Accreditation Status — https://www.fda.gov/about-fda/cdrh-transparency/510k-third-party-performance-metrics-and-accreditation-status 4. FDA, 510(k) Third Party Review Program guidance — https://www.fda.gov/regulatory-information/search-fda-guidance-documents/510k-third-party-review-program 5. FDA, Premarket Notification 510(k) — https://www.fda.gov/medical-devices/premarket-submissions-selecting-and-preparing-correct-submission/premarket-notification-510k 6. FDA, Medical Device User Fee Rates for Fiscal Year 2026, Federal Register — https://www.federalregister.gov/documents/2025/07/30/2025-14412
This article summarises publicly available FDA information and is intended as general regulatory awareness. It is not legal or regulatory advice.
Follow MedTech Insights
New articles on FDA 510(k) and De Novo pathways, CE Mark and EU MDR, and device reimbursement — sent to your inbox as they publish.
Comments
No comments yet. Be the first to add your read on this.